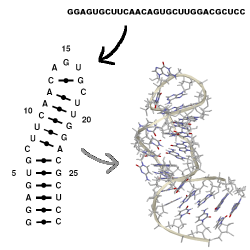
RNA secondary structure prediction from sequence, with non-canonical base pairs
MC-Fold help
MC-Fold predicts RNA secondary structures from sequence using nucleotide cyclic motifs (Parisien & Major, Nature 2008). Results are identical to the original MC-Fold web service.
Sequence
Up to 150 nucleotides. Upper or lower case is accepted; T is read as U.
Dashes, underscores and spaces are ignored. Modified-nucleotide symbols accepted by the original MC-Fold are also accepted.
Algorithm options
- H-type pseudoknots: also consider H-type pseudoknotted structures (slower).
- Return the best N structures: number of structures listed in the results.
- Explore the best X % sub-optimal structures: energy window explored around the optimum. Larger values are slower.
Structural constraints
A mask of the same length as the sequence: ( ) paired, . unpaired, * unconstrained.
Check “convert” to turn all . into *.
Reactivity data
Masks of the same length as the sequence marking reactive nucleotides with x. High, medium and low reactivity add
+8, +4 and +2 kcal/mol penalties respectively when the nucleotide is paired.
Results
Each job gets a private link that stays valid for 30 days. The results page lists the best structures with their energies (kcal/mol), a base-pair consensus, the best structure in CT format, and the structures found for each number of stems.
Scripted access
Links and scripts using the original address
/cgi-bin/MC-Fold/mcfold.static.cgi?sequence=…&top=…&explore=… keep working.